FDA Finalizes Cell And Gene Therapy Potency Guidance, Putting Manufacturing And Supply Controls in Focus
By Elizabeth Urbanek, Executive Editor
The U.S. Food and Drug Administration (FDA) has issued final guidance outlining how manufacturers of cellular and gene therapy products should demonstrate and maintain product potency throughout development and commercialization.
Released Nov. 2, the Potency Assurance for Cellular and Gene Therapy Products guidance replaces the FDA's January 2011 guidance on potency testing. It recommends a science- and risk-based approach that considers manufacturing process design, material controls, in-process testing, and lot-release assays.
The guidance applies across the product life cycle, from early IND submissions through biologics license application approval and post-approval monitoring. It also addresses considerations specific to viral vector therapies, patient-specific products, and therapies with short shelf lives.
For clinical supply teams, one part deserves particular attention: the FDA's recommendations extend beyond manufacturing and laboratory testing. The agency identifies storage, shipping, handling, container closures, and delivery devices as factors that can affect product potency. The guidance represents the FDA's current recommendations and does not establish new legally enforceable requirements for manufacturers.
Read the FDA's final guidance.
What This Means For Clinical Supply
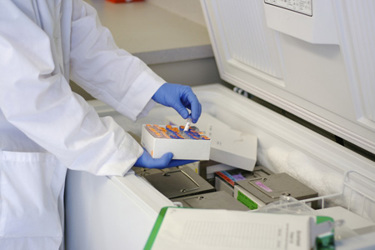
A product's potency can be affected by what happens after manufacturing is complete. Storage conditions, transportation, handling, and preparation before administration all matter. The FDA specifically identifies these areas in its guidance, along with container closures and delivery devices.
For clinical supply teams, that puts some familiar operational decisions into the potency discussion. How will the product be shipped? What temperature conditions must be maintained? How much time is available between release and administration? Are the treatment site and the people handling the product prepared for its arrival?
Those questions become more complicated with cell and gene therapies. Some products are made for individual patients, have short shelf lives, or require specialized temperature-controlled transportation. Manufacturing and release schedules have to line up with shipping arrangements and treatment-site availability. There isn't always much room to recover when something goes wrong.
The FDA also discusses the materials used to make these therapies. Starting materials and other manufacturing inputs can affect potency, so manufacturers need to understand their quality and account for variability. That includes evaluating suppliers and considering whether a change in materials or manufacturing processes could affect the finished product.
The same attention is needed as a therapy moves through clinical development. Stability data help sponsors establish how long a product can be stored and under what conditions. Those details influence how much time supply teams have to arrange transportation, coordinate with clinical sites, and get the product ready for administration.
For a patient-specific therapy, a missed shipment window or a delay in release testing can have immediate consequences for a scheduled treatment. Supply arrangements need to account for those possibilities before the first product is ready to ship, not after a problem occurs.
The FDA's guidance is directed at manufacturers, but the issues it raises reach well beyond the production facility. Clinical supply decisions about packaging, storage, transportation, and handling are part of getting a product to a patient without compromising its quality.
The Supply Question
As cell and gene therapy development continues, how early are sponsors bringing clinical supply teams into decisions that could affect product potency?
Additional Reading:
- Managing Operational Challenges In CAR-T Clinical Trial Logistics
- How Clinical Supply Is Shaping The Next Generation Of Cell And Gene Therapies
About The Author:
 Elizabeth Urbanek is executive editor of Clinical Supply Leader, where she works with clinical trial supply and logistics professionals across pharma and biotech to bring real-world perspectives, operational challenges, trends, scientific insights, and regulatory strategy to the field.
Before joining life sciences media, Elizabeth spent more than 15 years in broadcast, digital, and live-event storytelling, helping highly technical teams communicate complex ideas with clarity and impact. She brings that same approach to clinical supply, creating editorial space for experts to share what is happening across clinical trial operations.
Elizabeth is passionate about connecting people and ideas, identifying emerging themes, and showcasing work that can move the clinical supply field forward.